uconn health

Biosafety for Clinical Research

Conducting Clinical Research Involving Biological Materials at UConn Health

Overview

Clinical research at UConn Health that involves biohazardous human specimens, biological agents, recombinant or synthetic nucleic acids, viral vectors, genetically modified organisms, or live biotherapeutic products is reviewed by the Biosafety Program to ensure appropriate biosafety controls are in place.    Working with biohazardous materials may also necessitate Institutional Biosafety Committee (IBC) review prior to study initiation. This page outlines biosafety expectations, review pathways, and investigator responsibilities to ensure safe and compliant research conduct.

When to Consider Biosafety Program Review

  • Processing of biohazardous human specimens
  • Clinical administration of investigational biological products
  • Gene or viral vector therapies
  • Engineered cells or genetically modified organisms
  • Live microorganisms or microbiome therapeutics
  • Infectious agents or biological toxins

Biosafety Review Process
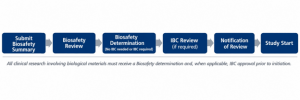

Investigator Responsibilities

  • Disclose all biological agents and materials during IRB study submission
  • Complete PI  Biosafety Summary (Every 2 Years when requested, or as research updates)
    Submit here
  • Obtain Biosafety or IBC authorization before initiating covered activities
  • Ensure congruency between clinical protocol, funding, and PI Biosafety Summary or registration.
  • Ensure biosafety compliance and staff training (see training requirements below)
  • Submit amendments for changes that have the potential to affect biological risk to the Biosafety Office using the PI Biosafety Summary.

Required Biosafety Program and Environmental Health and Safety Training

All personnel involved in clinical research involving biological materials must complete required training prior to study initiation and maintain training as applicable.

For Principal Investigators (PIs)

1. Initial Lab Safety (Chemical & Biosafety / Bloodborne Pathogens Training link

2. CITI PI Biosafety/Biosecurity Training (Every 3 Years)  CITI Program

Register an account and be sure to affiliate to University of Connecticut Health Center

We have a contract with CITI Program for online compliance training (https://about.citiprogram.org/en/homepage/ ). To access it you need to register (upper right of homepage) as an affiliate of “University of Connecticut Health Center” (and no other). Once you register and promise not to divulge the material in the trainings to others, you will need to navigate to the courses:

  1. When you come to “CITI Course Enrollment Procedure”
  2. Scroll all the way to the bottom under “question 8”
  3. Click the checkbox next to “PI” under Biosafety/Biosecurity
  4. Click the “Complete Registration” box
  5. Click the “Finalize Registration” box
  6. Click “view courses” in the upper part;
    1. Note: If you do not see courses as below, scroll down to “Learner Tools for University of Connecticut Health Center” and click “Add a course”. This will take you to the Select Curriculum page where you scroll all the way to the bottom where it says “Question 8”.
  7. Choose your learner group
  8. Click “Start Now”, agree to the Assurance statement, and take all modules in the course. There are quizzes


    3. DOT/IATA Shipping / Transport Training (If moving or shipping biological samples)
    Watch video

    4. Annual Bloodborne Pathogen refresher training ongoing through SABA, Healthstream, or the EH&S Training page (annually after Initial Lab Safety training)

    For Research Staff

    1. Initial Lab Safety (Chemical & Biosafety / Bloodborne Pathogens
      Training link
    2. Annual Bloodborne Pathogen refresher training ongoing through SABA, Healthstream, or the EH&S Training page (annually after Initial Lab Safety training)
    3. DOT/IATA Shipping / Transport Training (If applicable)
      Watch video

    Additional EH&S trainings may be required. Per UConn Health’s policy and OSHA’s Hazard Communication Standard, also known as “The Laboratory Standard” it is mandated that ALL people working in laboratories covered by the standard at UConn Health (including the vivarium and TIPS space) require training upon their initial assignment. This is in addition to the basic training they attend at orientation. The training in the safe use of chemicals, biosafety, emergency response and waste management is essential for a safe working environment.

    EH&S training website

    https://ovpr.uchc.edu/services/rics/ehs/safety-training-sessions/

    EH&S Contact Information:


    If Biosafety Review Was Missed

    If clinical research involving biological materials is identified without required BioSafety review, the UCH
    Biosafety Program will perform a risk assessment and coordinate corrective actions to achieve
    compliance. Approvals apply prospectively; retrospective approval cannot be granted.

    • IBC submission or amendment required
    • Potential activity pause for higher‑risk work
    • Reportability assessment under regulations/policy
    • Training or process improvements as needed


    Copilot language:

    Identification of Clinical Research Involving Biological Materials Without Required Biosafety Review

    If clinical research involving biological materials is identified without the required Biosafety review and approval, the UCH Biosafety Program will work collaboratively with the research team to assess potential risks and determine appropriate corrective and preventive actions (CAPA). Our primary goal is to ensure the safety of personnel, participants, and the environment while helping investigators achieve compliance moving forward.

    Because Biosafety approvals can only be granted prospectively, retrospective approval cannot be provided for activities that have already occurred. However, in most cases, the review process will focus on understanding the circumstances, addressing any gaps, and implementing appropriate measures to prevent recurrence.

    Potential outcomes may include:

    • Submission of an IBC application or amendment to document and review the activity.
    • A risk-based assessment to determine whether any research activities should be modified or temporarily paused while review is completed.
    • Evaluation of whether reporting obligations exist under applicable regulations, institutional policies, or sponsor requirements.
    • Additional training, process improvements, or other corrective actions to support ongoing compliance.

    The Biosafety Program is committed to partnering with investigators to address issues proactively and promote a culture of safety and compliance.


    Contact Biosafety

    Contact the Biosafety Program Office with any questions or for more information: IBC@uchc.edu.

    Welcome to the Office of Clinical & Translational Research (draft)

    The Office of Clinical & Translational Research (OCTR) supports investigators, research staff, and sponsors by providing expert guidance through the complexities of clinical trial budgets, contract negotiations, and financial compliance at UConn Health.

    Our Mission

    We ensure that all clinical research conducted at UConn Health complies with federal and state regulations as well as institutional policies — while helping researchers move their studies forward efficiently and responsibly.

    What We Do

    Our team partners with researchers to provide:

    Budget Development & Management

    • Build detailed Budget Workbooks for each study to compile an internal study cost estimate.
    • Collaborate with PIs to create viable budgets for all clinical trials opened at UConn Health that accrue that accrues John Dempsey Hospital (JDH) and/or UConn Medical Group (UMG) and/or dental charges.
    • Establish and reconcile Banner research accounts

    Contract Negotiation

    • Industry-sponsored and investigator-initiated trials
    • University-to-university agreements
    • Cooperative group and foundation-supported trials

    Financial Compliance & Oversight

    • Audit ongoing clinical trials for compliance with institutional, state, and federal requirements

    Connect with Us

    Getting Started (draft)

    OCTR provides resources and guidance to help researchers and study teams navigate the clinical trial process at UConn Health. This page provides an overview of the steps involved in getting started, along with key procedures, definitions, and commonly used acronyms. The information provided is intended to help study teams understand the OCTR process and identify the appropriate resources throughout the life of a clinical trial.

    Budget & Contract Procedure

    Budget Process: Determination Schema for Industry Sponsored, Industry Supported, University to University, Co-operative Group or Foundation Supported Clinical Trials

    A. PRE-TRIAL:

      • In order to open a clinical trial at the University of Connecticut Health Center, the following question must be answered to determine the proposal’s flow through the institution:
        • Does the project involve human subjects?
      • All studies require a prelim packet so a budget and memo can be provided to the study team. At this time, the Budget & Financial Specialist will determine if the study only requires an expedited memo – this is required for IRB submission.

    B. BUDGET WORKBOOK:

    • OCTR staff complete the Budget Workbook once the prelim packet is received from the study team:
      • Items and services are delineated as Protocol Induced Costs (PIC) or Routine Clinical Care (RC)
      • T&E is designated for dedicated research staff
    • Once the PI receives, reviews, and approves the budget workbook they provide the Budget & Financial Specialist with the approval to proceed with negotiations.
    • Budget & Financial Specialist commences budget negotiations and/or PI provides additional funding sources.

    C. INITIATION OF ROUTING:

    • Upon OCTR completion of full contract negotiations (budget and contract), the PI approves the budget and signs the Internal Proposal Review Form (IPR), thereby initiating the routing process. Routing sheet is then signed by:
      • PI’s Department Chair
      • Co-Investigator (if applicable)
      • Co-Investigator’s Department chair
      • Dean
      • Director, Sponsored Program Services

    D. CONTRACT

    Contract negotiations continue in parallel with the budget preparation and the routing, as shown on the preceding flow chart above.

    • Contract negotiations may be ongoing at the time of initial IRB submission.

    E. IRB FINAL PROCESS

    • Study team may now submit a complete final application packet to the IRB per institutional guidelines, with the memo certifying that a Budget Workbook is complete from Budget & Financial Specialist and the fully executed agreement from the Contract Specialist.

    F. POST-IRB APPROVAL PROCESS

    • Study cannot open until there is a fully executed contract and final IRB approval.
    • Upon final IRB approval, Budget & Financial Specialist will reach out to schedule the budget initiation meeting.
    • Budget initiation meeting occurs.
    • PI signed document received by OCTR.
    • Study Opens

    Definitions

    Budget Workbook

    Electronic tool used by OCTR in conjunction with study team to identify, segregate and monitor charges

    Clinical Research

    1. Patient-oriented research. Research conducted with human subjects (or on material of human origin such as tissues, specimens and cognitive phenomena) for which an investigator (or colleague) directly interacts with human subjects. This area of research includes:
    • Mechanisms of human disease
    • Therapeutic interventions
    • Clinical trials
    • Development of new technologies
    1. Epidemiologic and behavioral studies.
    2. Outcomes research and health services research.

    Clinical Trial

    A clinical trial is a systematic, organized, prospective intervention study in human subjects that is conducted according to a formal study plan (protocol) and that has measurable efficacy and/or safety-related outcomes that are amenable to statistical analysis. It employs one or more intervention technique(s) including prophylactic, screening, diagnostic, or therapeutic agents, devices, or procedures. It must have the approval of the IRB or the IRB’s review with a determination of exemption. Clinical trials are distinguished from other types of clinical research (e.g., behavioral research) that may need IRB approval but do not meet the other criteria of clinical trials.

    Clinical Trials Initiation Form

    Upon IRB approval, this form is generated by OCTR  to notify all departments involved of the services to be charged to the trial. This form includes critical billing information such as: BANNER account number,  party responsible for EPIC Research Billing Review, name of recipient of Transfer Vouchers, list of services to be charged, and category of sponsor.

    Preliminary Budget Information Packet

    OCTR document that initiates the Budget Workbook process. Packet is emailed to PI/Study Coordinator in response to query regarding doing a Budget Workbook; it is completed by the study team and returned to OCTR via email prior to study team meeting with OCTR staff

    Protocol Induced Costs (PIC)

    Items and services that are specifically excluded from the definition of Routine Clinical Services in a qualifying clinical trial such as:

    • The investigational item or service itself
    • Items and services provided solely to satisfy data collection and analysis needs and that are not used in the direct clinical management of the patient
    • Items and services customarily provided by the research free of charge for any enrollee in the trial
    • Items and services provided solely to determine trial eligibility

    Qualifying Clinical Trial

    As per the Medicare National Coverage Determination (“Routine Costs in Clinical Trials, also known as the CMS Clinical Trial Policy or “CMS CTP”): Any clinical trial receiving Medicare coverage of routine costs must meet both the basic three requirements and seven desirable characteristics to qualify for coverage.

    https://www.cms.gov/Regulations-and-Guidance/Guidance/Transmittals/downloads/R74NCD.pdf

    “Deemed” studies automatically meet the criteria for the seven desirable characteristics.

    Routine Clinical Services (RC)

    Routine costs of a clinical trial include all items and services that otherwise would be generally available to Medicare beneficiaries regardless of whether they are assigned to the experimental or control arm of a qualifying clinical trial. “Generally available” means that the item or service falls within an existing Medicare benefit category that is not excluded from coverage by another provision in the law and that is not the subject of an NCD stating that the item or service is not covered. If an item is covered outside of a clinical trial it is covered within the clinical trial.

    Study

    In this context, synomous with clinical trial/clinical research

    Acronyms

    CDA                Confidential Disclosure Agreement 

    CPT                 Clinical Procedural Terminology. codes for billing purposes 

    CTA                  Clinical Trial Agreement 

    CRC                Clinical Research Center 

    DCRC             Dental Clinical Research Center 

    F & A               Facilities & Administrative costs (a.k.a. indirect costs) 

    FOAPAL         Fund-Organization-Account-Program-Activity-Location. The abbreviation for  the chart of accounts structure used by the Finance in the BANNER system 

    IDC                 Indirect Costs (a.k.a. OH) 

    IP                     Intellectual Property 

    IPR                  Internal Proposal Review Form 

    IRB                  Institutional Review Board 

    JDH                 John Dempsey Hospital 

    MCA                Medicare Coverage Analysis 

    NCD                National Coverage Decision (Medicare) 

    NCI                  National Cancer Institute 

    NIH                  National Institutes of Health; “NIH” 

    OCTR              Office of Clinical and Translational Research 

    OH                   Overhead (a.k.a. IDC) 

    PI                      Principal Investigator 

    PIC                  Protocol Induced Costs 

    RC                    Routine Care (a.k.a. SOC) 

    SC                    Study Coordinator 

    SOC                Standard of Care (a.k.a. RC)  

    SOM                School of Medicine 

    SODM             School of Dental Medicine 

    SPS                  Sponsored Program Services 

    T & E                Time and Effort 

    TV                     Transfer Voucher 

    UMG                UConn Medical Group 

    Contracts & Budgets (draft)

    Process Overview

    All clinical trials require a confidential disclosure agreement (CDA) and a clinical trial agreement (CTA) that includes a budget with payment terms.

    The contract specialist works on the CDA first. Once the CDA is complete, the legal terms of the CTA are negotiated by the contract specialist and the budget and payment terms of the CTA is negotiated by the budget specialist in tandem.

    Contracts Information and Negotiation Process

    The OCTR negotiates industry-sponsored or investigator-initiated industry-supported contracts, university-to-university agreements, co-operative group contracts and foundation-supported contracts, which include the following:

    • Clinical Trial Agreements (CTA)
    • Master Clinical Trial Agreements (MCTA)
    • Confidentiality Disclosure Agreements (CDA)
    • Contract Amendments
    • Letters of Indemnification (LOI)

    For any other contract types, please reach out to SPS.

    Contract Negotiation Process

    The contract negotiation process is dependent on the type of agreement:

    Industry Sponsored Clinical Trial Agreements

    • The sponsor will initially send a non-disclosure agreement (NDA) or a confidentiality disclosure agreement (CDA) to the researcher. Once received, the researcher should forward the CDA to the contract specialist in the OCTR. The CDA will then be reviewed, negotiated, approved, and executed. Thereafter, the researcher will review the data and the study components in order to determine whether to participate in the study.
    • After reviewing the sponsor’s data and proposed study, the researcher and the sponsor will make a mutual decision whether or not to engage in the study. If the researcher decides to engage in the study, the sponsor will send a clinical trial agreement (CTA) to the researcher or the researcher’s study coordinator. The clinical trial agreement is then forwarded to the contract specialist in the OCTR for review and negotiation. Once the CTA is negotiated, the contract specialist will obtain signatures from an authorized representative of UConn Health, the researcher, and the sponsor.

    University to University Agreements

    • University to University agreements are subcontracts with a university that has a governing contract with a company sponsor. The negotiation procedure for University to University agreements is similar to the negotiation procedure for industry-sponsored clinical trial agreements.

    Investigator-Initiated Clinical Trial Agreements

    • An investigator-initiated clinical trial is one that is authored by the investigator and financially supported by industry, a foundation, or another university. The funding agency may send its own company contract to OCTR for negotiation, or OCTR may send the funding agency a clinical trial agreement generated by UConn Health. Thereafter, the negotiation procedure is generally tantamount to the process described above for industry-sponsored studies.

    Cooperative Group Studies

    • UConn Health has master agreements with cooperative groups which are designed to promote and support clinical trials. OCTR negotiates all agreements pertaining to the cooperative group studies or its corresponding master contract.

    Budget Information and Negotiation Process

    The budget development and negotiation process generally includes the following: 

    • The sponsor or funding agency provides the proposed budget and/or budget template to the researcher/study team. The budget should be forwarded to OCTR for review and negotiation (octrclinicaltrial@uchc.edu). 
    • OCTR performs a Medicare coverage analysis by reviewing the protocol, schedule of events, and creates an internal proposed budget to identify the costs associated with conducting the study at UConn Health. 
    • OCTR works with the study team and applicable UConn Health department(s) to obtain appropriate internal costs for study-related services and resources. 
    • OCTR develops and negotiates the study budget to ensure that clinical procedures, laboratory testing, pharmacy services, imaging, pathology, personnel effort, administrative activities, and other study-specific expenses are appropriately reflected in the budget. 
    • OCTR negotiates the proposed budget with the sponsor or funding agency and works with the study team to address questions or requested revisions. 
    • Once the budget has been negotiated and finalized, OCTR coordinates with the contract specialist, as necessary, to ensure that the final budget is consistent with the terms of the applicable agreement. 
    • Budget amendments may be required when changes to the protocol, schedule of events, study procedures, personnel, internal costs, or other study-related requirements result in additional costs. OCTR reviews and negotiates applicable budget changes to ensure that the revised budget appropriately reflects the costs of conducting the study. 

    Our Mission

    The Human Research Protection (HRP) Education & Outreach (E&O) program is a unified cross-campus initiative to provide education and outreach to all who are involved in research involving human subjects. Our mission is to equip researchers with the knowledge and tools needed to navigate the regulatory landscape of human subjects research.

    Through proactive engagement, the program aims to support researchers and promote ethical practices that enhance understanding of human subjects protections across the university community.

    The HRP Education & Outreach program goals are to:

    • Inform the research community about the latest developments in human subjects protection, including news, guidance, and regulatory updates.
    • Educate researchers, IRB members, and IRB staff through tailored outreach and ongoing learning opportunities.
    • Support investigators through compliance reviews, self-monitoring tools, and personalized assistance to promote high-quality, ethically sound research.

    By offering these resources, we seek to reduce compliance burdens and foster a culture of collaboration and integrity in research involving human participants.

    Contact Us

    We are here to support your research. Reach out to ask questions, request a consultation, or share feedback.

    Email: hrp-education@uconn.edu

    Team Contacts:

    • Joan Levine, MPH, CIP – Team Lead
    • Rebecca Burke, MS
    • Ellen Ciesielski

    Compliance Monitoring

    *New* PI Self-Assessment

    The Education & Outreach unit is introducing a new PI Self-Assessment process this fall, as a voluntary component of the post-approval monitoring program. The new PI Self-Assessment serves as an alternative approach for certain studies, offering a more flexible and educational approach to research oversight. This option will be available for studies that are classified as minimal risk by the IRB, excluding studies subject to FDA regulations. Completing a self-assessment removes you from the routine monitoring process for minimal risk research for a period of 2 years.


    If your study is selected for a self-assessment, you will receive a PI Self-Assessment Form and Record Review Form from the Education & Outreach (E&O) team. If you agree to participate, complete and return the form within 30 days of notification. After submission, the E&O team may follow up with questions, request a brief meeting for clarification, or review applicable documents. As a final step, an audit summary report will be sent to you for your records.


    Studies not eligible for this process will continue to receive traditional routine post-approval monitoring.

    Human Research Protection Compliance Monitoring Program

    Human subjects research at UConn and UConn Health is monitored by the HRP Education & Outreach program. The goal of the monitoring program is to assess compliance of human subjects research with federal, state, local law, and UConn policies, identifying areas for improvement, and providing recommendations and support based on best practices, current policies and the principles set forth in the Belmont Report.

    Studies may be randomly selected for a compliance review or chosen for other reasons as described in the applicable policies and procedures. Categories of compliance reviews include routine, self-assessment, informed consent, for-cause, IRB-directed, and investigator-initiated. All active studies are subject to such reviews, including exempt research or studies where a reliance agreement is in place with another institution.

    A detailed report summarizing the review findings, required actions, and recommendations will be sent to the researcher. Additionally, findings from the review may be shared with individuals responsible for research oversight to reinforce compliance with policies and regulatory requirements. Findings from these reviews help inform and shape future educational offerings. More information about the monitoring program can be found in our policies and standard operating procedures.

    Related Policies & Procedures:

    Institutional Review Entity (IRE)

    The Institutional Review Entity (IRE) is established by the research institution to fulfill oversight responsibilities as outlined in USG DURC-PEPP Policy and the USG Implementation Guidance for identification, review, and oversight of life sciences research that is within Category 1 and Category 2 as defined by that policy.

    On May 5, 2025, a new Executive Order (EO) was issued calling for modifications to the 2024 United States Government Policy for Oversight of Dual Use Research of Concern and Pathogens with Enhanced Pandemic Potential (often referred to as the “USG DURC-PEPP Policy”).

    As the institution awaits additional guidance on how the provisions outlined in the May 5, 2025, EO may impact future research operations, researchers should continue to adhere to all applicable rules, regulations, and processes under the purview of UCH IRE, including all expectations set forth by the current USG-DURC-PEPP Policy (outlined in greater detail below).

    UCH implements these federal oversight requirements through the UCH Life Sciences Research of Concern Policy, which outlines institutional responsibilities, investigator expectations, review procedures, and risk mitigation requirements for research falling within the scope of the USG DURC-PEPP Policy.

    About the USG DURC-PEPP Policy

    At UCH, the USG DURC-PEPP Policy applies to all research, regardless of funding source, which meets the criteria for DURC-PEPP research. Federal sponsors may delay the release of funds if a project identified as DURC-PEPP research does not fulfill all requirements for compliance with the USG DURC-PEPP Policy. This includes research funded or sponsored by federal grants, contracts, cooperative agreements, and other agreements. The United States Government  USG DURC-PEPP Policy supersedes previous DURC policies and the 2017 Enhanced Potential Pandemic Pathogens Framework (P3CO). The USG DURC-PEPP Policy does not supersede, but complements, other existing federal regulations, including the Select Agent Regulations.

    All UCH Principal Investigators (PIs) proposing or conducting research involving biological agents and toxins must assess whether or not their research potentially falls under the USG DURC-PEPP Policy.  Specifically, PIs proposing to work with or generate any replication-competent infectious agent or proposing to work with a toxin of any amount from the Federal Select Agents and Toxins list must make an assessment of whether the research is reasonably anticipated to be within the scope of Category 1 or Category 2 research. If DURC-PEPP research is identified, the PI must work with the UCH Institutional Review Entity (IRE) (ibc@uchc.edu) to develop a risk-benefit assessment and risk mitigation plan that the funding agency must approve before the work can begin.


    Categories of Research


    Category 1 Research


    Scope: Involves biological agents or toxins from a pre-determined list, which includes over 80 specific agents and toxins (including all select agents and toxins + all RG4 and many/most RG3 agents). These include Bacillus cereus Biovar anthracis, Botulinum neurotoxins, Coxiella burnetii, Ebola virus, Francisella tularensis, SARS-CoV, and others.


    Experimental Outcomes: The research is reasonably anticipated to result in, or does result in, one or more of the following nine outcomes:


    Increase transmissibility of a pathogen within or between host species.

    Increase the virulence of a pathogen or convey virulence to a non-pathogen.

    Increase the toxicity of a known toxin or produce a novel toxin.

    Increase the stability of a pathogen or toxin in the environment or increase its ability to disseminate.

    Alter the host range or tropism of a pathogen or toxin.

    Decrease the ability to detect a human or veterinary pathogen or toxin using standard diagnostic or analytical methods.

    Increase resistance of a pathogen or toxin to clinical or veterinary prophylactic or therapeutic interventions.

    Alter a human or veterinary pathogen or toxin to disrupt the effectiveness of preexisting immunity via immunization or natural infection.

    9.Enhance the susceptibility of a host population to a pathogen or toxin.


    Category 2 Research


    Scope: Involves a Pathogen with Pandemic Potential (PPP) or a Pathogen with Enhanced Pandemic Potential (PEPP). These include pathogens like H5N1 influenza virus, SARS-CoV-2, and MERS-CoV.


    Experimental Outcomes: The research is reasonably anticipated to result in, or does result in, one or more of the following four outcomes:

    Enhance transmissibility of the pathogen in humans.

    Enhance the virulence of the pathogen in humans.

    Enhance the immune evasion of the pathogen in humans by modifying it to disrupt pre-existing immunity via immunization or natural infection.

    Generate, use, reconstitute, or transfer an eradicated or extinct PPP or a previously identified PEPP.


    Responsibilities


    Researchers/PIs:

    Conduct initial and ongoing self-assessments of research associated with the agents that fall under the DURC policy.

    Collaborate with the IRE on risk-benefit analyses.

    Draft and implement Risk Mitigation Plans (RMPs) in coordination with the IRE when research is classified as Category 1 or 2.

    Ensure ongoing compliance with approved RMPs for Category 1 or 2 research.


    Institutional Oversight:

    Establish and maintain internal policies/procedures for DURC and PEPP oversight.

    Maintain records of assessments, RMPs, and compliance activities.

    Assess research with agents that fall under the new DURC policy to determine if research falls under Category 1 or 2.

    Submit required reports to federal agencies.

    Provide education and training to researchers and staff involved in high-risk research.


    Appendix: Category 1 Agents and Toxins List:

    HHS Select Agents and Toxins 


    Abrin

    Bacillus cereus Biovar anthracis 

    Botulinum neurotoxins

    Botulinum neurotoxin producing species of Clostridium 

    Conotoxins (Short, paralytic alpha conotoxins containing the following amino acid sequence X1CCX2PACGX3X4X5X6CX7)

    Coxiella burnetii

    Crimean-Congo haemorrhagic fever virus

    Diacetoxyscirpenol

    Eastern Equine Encephalitis virus

    Ebola virus

    Francisella tularensis

    Lassa fever virus

    Lujo virus

    Marburg virus

    Mpox virus

    Reconstructed replication competent forms of the 1918 pandemic influenza virus containing any portion of the coding regions of all eight gene segments (Reconstructed 1918 Influenza virus)

    Ricin

    Rickettsia prowazekii

    SARS-associated coronavirus (SARS-CoV)

    SARS-CoV/SARS-CoV-2 chimeric viruses resulting from any deliberate manipulation of SARS-CoV-2 to incorporate nucleic acids coding for SARS-CoV virulence factors

    Saxitoxin


    South American Haemorrhagic Fever viruses: 


    Chapare

    Guanarito

    Junín

    Machupo

    Sabia

    Staphylococcal enterotoxins (subtypes A,B,C,D,E)

    T-2 toxin

    Tetrodotoxin


    Tick-borne encephalitis complex (flavi) viruses:


    Far Eastern subtype

    Siberian subtype

    Kyasanur Forest disease virus

    Omsk hemorrhagic fever virus

    Variola major virus (Smallpox virus)

    Variola minor virus (Alastrim)

    Yersinia pestis


    Overlap Select Agents and Toxins 


    Bacillus anthracis

    Bacillus anthracis Pasteur strain

    Brucella abortus

    Brucella melitensis

    Brucella suis

    Burkholderia mallei

    Burkholderia pseudomallei

    Hendra virus

    Nipah virus

    Rift Valley fever virus

    Venezuelan equine encephalitis virus


    USDA Veterinary Services (VS) Select Agents and Toxins 


    African horse sickness virus

    African swine fever virus

    Avian influenza virus

    Classical swine fever virus

    Foot-and-mouth disease virus

    Goat pox virus

    Lumpy skin disease virus

    Mycoplasma capricolum 

    Mycoplasma mycoides

    Newcastle disease virus

    Peste des petits ruminants virus

    Rinderpest virus

    Sheep pox virus

    Swine vesicular disease virus


    USDA Plant Protection And Quarantine (PPQ) Select Agents and Toxins 


    Coniothyrium glycines (formerly Phoma glycinicola and Pyrenochaeta glycines)

    Peronosclerospora philippinensis (Peronosclerospora sacchari)

    Ralstonia solanacearum

    Rathayibacter toxicus

    Sclerophthora rayssiae

    Synchytrium endobioticum

    Xanthomonas oryzae


    For biological agents affecting humans that have not been assigned a Risk Group in the NIH Guidelines, agents affecting humans that are recommended to be handled at BSL3 or BSL4 per the BMBL guidance are subject to the USG DURC-PEPP Policy


    NIH Guidelines for Research Involving Recombinant or Synthetic Nucleic Acid Molecules (NIH Guidelines), Appendix B, Risk Group 4 and subset of Risk Group 3 


    Risk Group 4 (RG4) – Viral Agents 


    Arenaviruses

    Guanarito virus

    Lassa virus

    Junin virus (except the candid #1 vaccine strain listed in Appendix B-II-D Risk Group2 (RG2) – Viruses)

    Machupo virus

    Sabia

    Bunyaviruses (Nairovirus)

    Crimean-Congo hemorrhagic fever virus

    Filoviruses

    Ebola viruses

    Marburg viruses

    Flaviruses – Group B Arboviruses

    Tick-borne encephalitis virus complex including Absetterov, Central European encephalitis, Hanzalova, Hypr, Kumlinge, Kyasanur Forest disease, Omsk hemorrhagic fever, and Russian spring-summer encephalitis viruses

    Herpesviruses (alpha)

    Herpesvirus simiae (Herpes B or Monkey B virus)


    Paramyxoviruses

    Equine Morbillivirus (Hendra virus)

    Hemorrhagic fever viruses as yet undefined


    Risk Group 3 (RG3) – Bacterial Agents Including Rickettsia* 


    Bartonella

    Brucella including B. abortus, B. canis, B. suis

    Burkholderia (Pseudomonas) mallei, B. pseudomallei

    Coxiella burnetii (except the Phase II, Nine Mile strain listed in Appendix B-II-A, Risk Group 2 (RG2) – Bacterial Agents Including Chlamydia)

    Francisella tularensis (except those strains listed in Appendix B-II-A, Risk Group 2 (RG2) – Bacterial Agents Including Chlamydia)

    Orientia tsutsugamushi (was R. tsutsugamushi)

    Pasteurella multocida type B -“buffalo” and other virulent strains

    Rickettsia akari, R. australis, R. canada, R. conorii, R. prowazekii, R. rickettsii, R. siberica, R. typhi (R. mooseri)

    Yersinia pestis (except those strains listed in Appendix B-II-A, Risk Group 2 (RG2) – Bacterial Agents Including Chlamydia)


    Risk Group 3 (RG3) – Viruses and Prions* 


    Alphaviruses (Togaviruses) – Group A Arboviruses

    Chikungunya virus (except the vaccine strain 181/25 listed in Appendix B-II-D Risk Group2 (RG2) – Viruses)

    Semliki Forest virus

    Venezuelan equine encephalomyelitis virus (except the vaccine strains TC-83 and V3526, see Appendix-II-D (RG2) – Viruses)

    Other viruses as listed in the reference source (see Section V-C, Footnotes and References of Sections I through IV)

    Arenaviruses

    Flexal

    Lymphocytic choriomeningitis virus (LCM) (neurotropic strains)

    Bunyaviruses

    Hantaviruses including Hantaan virus

    Rift Valley fever virus

    Coronaviruses

    SARS-associated coronavirus (SARS-CoV)

    Middle East respiratory syndrome coronavirus (MERS-CoV)

    Flaviviruses- Group B Arboviruses

    Japanese encephalitis virus (except those strains listed in Appendix B-II-D Risk Group2 (RG2) – Viruses)

    Yellow fever virus

    Other viruses as listed in the reference source (see Section V-C, Footnotes and References of Sections I through IV)

    Orthomyxoviruses

    Influenza viruses 1918-1919 H1N1 (1918 H1N1), human H2N2 (1957-1968), and highly pathogenic avian influenza H5N1 strains within the Goose/Guangdong/96-like H5 lineage (HPAI H5N1).

    Poxviruses

    Monkeypox virus (Clade I & Clade II containing nucleic acids coding for clade I MPVX virus virulence factors)

    Prions

    Transmissible spongiform encephalopathies (TSE) agents (Creutzfeldt-Jacob disease and kuru agents)(see Section V-C, Footnotes and References of Sections I through IV, for containment instruction)


    EXCLUDED RG3 Agents: 


    Human immunodeficiency virus (HIV) types 1 and 2

    Human T cell lymphotropic virus (HTLV) types 1 and 2

    Simian immunodeficiency virus (SIV)

    Mycobacterium tuberculosis, Mycobacterium bovis

    Clade II of MPVX viruses unless containing nucleic acids coding for clade I MPVX virus virulence factors

    Vesicular stomatitis virus

    Coccidioides immitis (sporulating cultures; contaminated soil)

    Histoplasma capsulatum, H. capsulatum var. duboisii

    Compliance

    Sponsored Program Services (SPS) is developing a comprehensive compliance program. As the program develops, opportunities will be listed on the links available on the sidebar.

    Guidelines – Stop Work Orders

    Federal Stop-Work or Grant Termination Directives

    These guidelines are intended to assist investigators who may be at risk for or have received a directive from a federal funding agency to stop, pause, terminate or otherwise prematurely end a human research study.  Please contact the IRB Office if you have any questions or need assistance.


    Euthanasia of Sick Animals by CCM Veterinary Staff

    Purpose

    In order to comply with federal regulations (USDA and PHS) and the Guide for the Care and Use of Laboratory Animals, UConn Health’s IACUC has established a policy on euthanasia of sick animals by CCM veterinary staff.

    Action

    1. When sick animals are found by animal caretakers (ACT), a sick report is written up and an email is sent to the PI and/or contact individual identified on cage cards.
    2. The CCM Veterinary Services is also cc’d on the e-mail correspondence for their evaluation.
    3. If the animal is displaying signs of humane endpoints as identified in the animal care and use protocol or has a Body Condition Score (BCS) of 2 or less, veterinary staff will send an email to the PI and/or contact individual with recommendation of euthanasia. This may or may not be after consultation with the Attending Veterinarian.  If the recommendation is for euthanasia of the animals, the email will include a due date and time for corrective action to the research staff. The PI must respond by email within 24 hours.  If no response is received, this will be interpreted as consent by the PI to the medical opinion of the veterinary staff as to the disposition of the animal.  The PI will be charged accordingly.
    4. If the veterinary staff identify the same animal during their routine veterinary clinical rounds, they will euthanize the animal and report it to PI and the IACUC.
    5. If the animal is moribund or in severe distress, the Veterinary Services can euthanize the animal immediately and notify the PI/research staff. If the animals are under Death as an Endpoint or EAE studies, they should be posted in the room or identified at cage level to avoid confusion.

    Effective Dates: January 31, 2025 through December 31, 2027

    This policy has been approved by a majority vote of the UConn Health IACUC